
The central problem
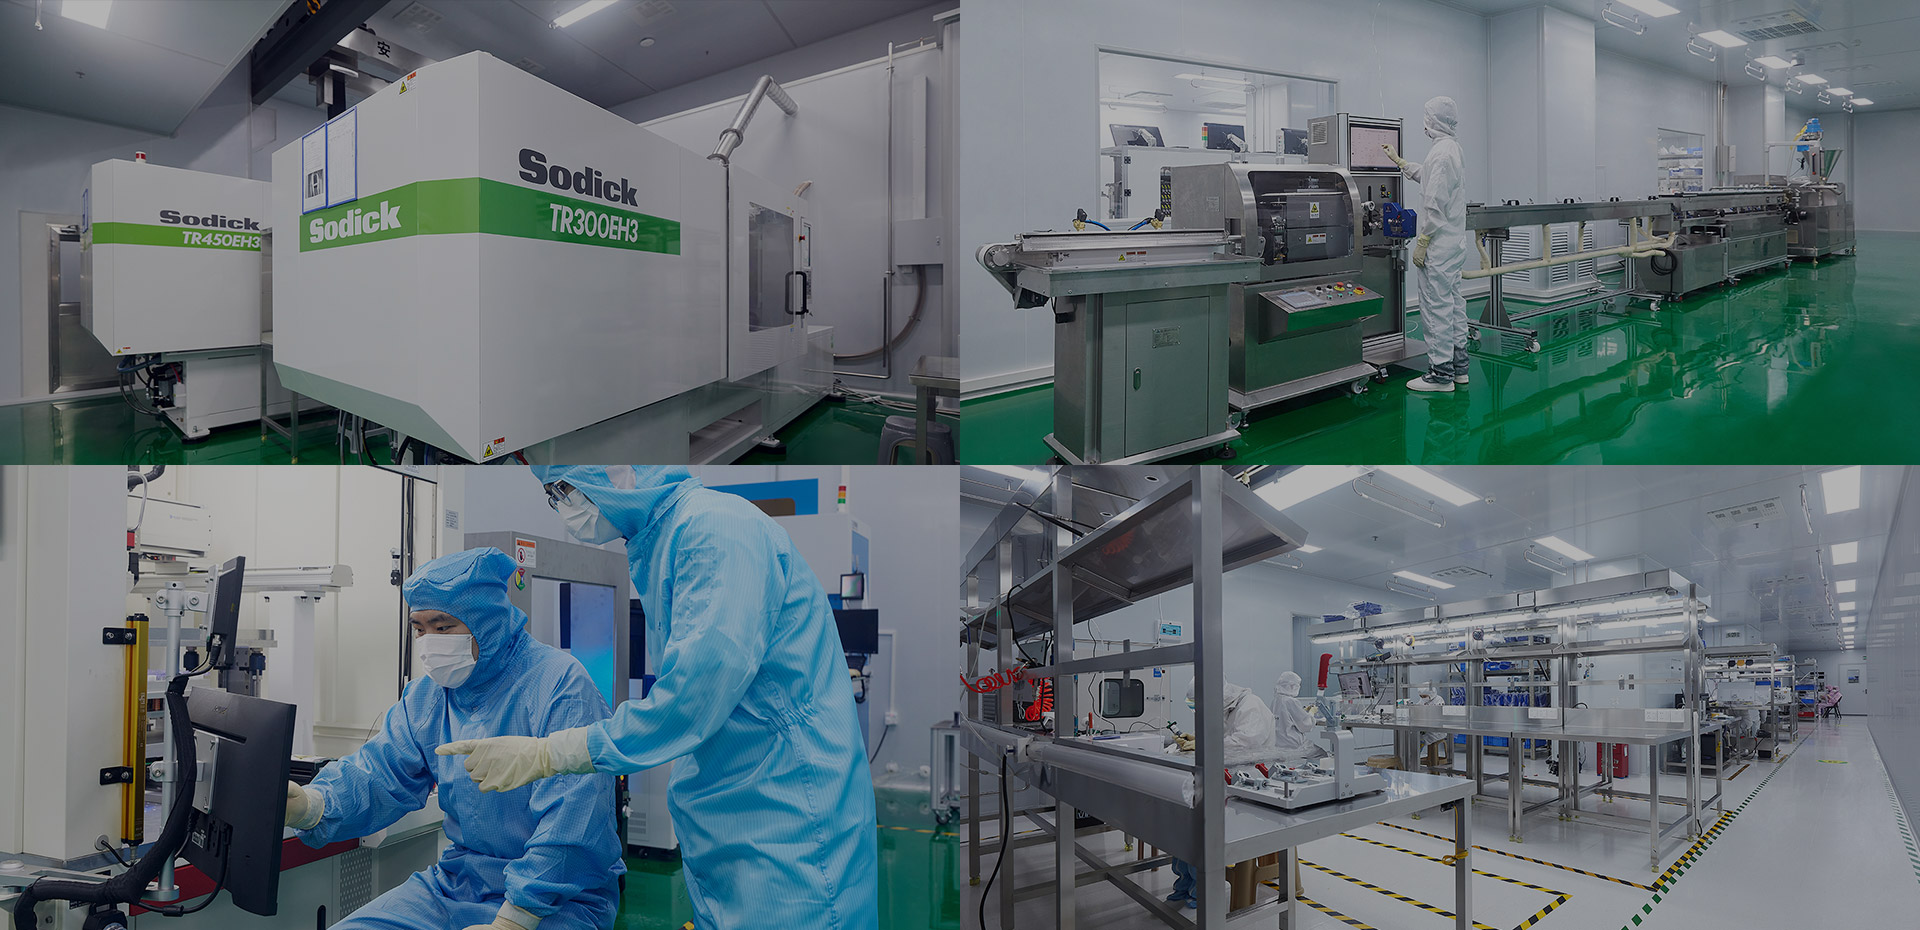
Manufacturers and design teams commonly misjudge what’s required to move from prototype to reliable production: consistent yields, traceable processes, and compliant test records. That misjudgment becomes visible at the medical device assembly line when poor tolerances, undocumented changes, or supplier bottlenecks force redesigns, regulatory delays, and shipment rework. The core issue isn’t only cost; it’s unknown risk that surfaces only after units leave QC.
How risk shows up in real projects
Losses appear as mismatched component lots, unclear change control, and unexpected cleanroom limits. Teams often accept a vendor’s capability statement without verifying sample-to-scale equivalence. A prototype that functions in a lab can fail at volume if tooling, fixturing, and inline inspection were not evaluated. Expect problems where contracts focus on price and not on documented process windows, acceptance criteria, or contingency plans for nonconforming lots.
EEAT: credentials, evidence, and a real-world anchor
Look for demonstrable experience: on-site engineers who have executed device transfers, quality managers with ISO 13485 audit histories, and published validation reports for sterilization and biocompatibility. Practical evidence includes first-article inspection reports, capability studies, and CAPA records you can review. Expertise is best confirmed by examples tied to recognized forums—work presented at the MicroTAS conference shows practical microfabrication and fluid handling experience relevant to devices that use microfluidic features. The same partners who can design controlled tooling and inline inspection for a microfluidic channel typically handle seal integrity and reagent handling with fewer surprises.
Technical checks that matter
Ask for hard proof: process capability (Cpk) for critical dimensions, traceable component lot histories, environmental monitoring logs for cleanrooms, and sample inspection data from runs at planned production volume. Verify tooling ownership, maintenance schedules, and first-article results under the device’s actual assembly conditions. Confirm test fixtures, software for automated inspection, and a clear change-control workflow that ties engineering change notices to documented risk assessments.
Alternatives and trade-offs
Choosing between a local partner, a Chinese contract manufacturer, or a hybrid strategy has concrete trade-offs. Local partners shorten travel and speed audits but can cost more per unit; Chinese manufacturers can offer scale and lower unit cost but require investment in remote oversight, stronger contractual protections, and staged transfers. A hybrid model—initial validation locally, volume production in China—reduces front-end risk but adds logistics complexity. Evaluate which trade-offs you can absorb without jeopardizing regulatory timelines.
Common mistakes and how to avoid them
Don’t accept generic certifications as proof of capability; request site-specific audit results tied to your product class. Avoid vague acceptance criteria—define dimensional limits, functional pass/fail thresholds, and allowable defect rates. Don’t skip a pilot run at target volumes. Protect IP with compartmentalized orders and documented data security procedures. Insist on routine reporting dashboards so you detect drift early rather than discover it at final inspection.
Conclusion: resolving the problem with measured partnership
Solving the production-choice problem requires precise checks, verified evidence, and a partner who documents process fidelity from tooling through lot release. When those elements align, risk drops and timelines shorten. For teams that need documented capability, process controls, and traceable production histories integrated with design support, kingsin represents a model of those elements working together rather than a single cost line on a spreadsheet.